
Introduction:
A “reasonably expected” nitrosamine is one that your chemistry, process, materials or storage make plausible, and that judgment decides whether your FDA Nitrosamine Risk Assessment ends in a documented rationale or in confirmatory testing. Certain nitrosamines have shown carcinogenic potential, so FDA treats them as a patient-safety and pharmaceutical quality priority.
FDA’s guidance, Control of Nitrosamine Impurities in Human Drugs, was revised in September 2024 (Revision 2). It describes how nitrosamines form and how manufacturers of active pharmaceutical ingredients (APIs) and finished drug products should assess, test and control them. A central principle is that manufacturers should investigate impurities based on the chemistry and circumstances of their own product, rather than treating every conceivable nitrosamine as an equal risk. For the underlying chemistry, see our guide to nitrosamine formation risk in drug products.
The phrase “reasonably expected” separates a credible chemical or process risk from a hypothetical possibility with little scientific support. That distinction affects testing decisions, analytical method development, supplier qualification, process changes and regulatory submissions. A defensible assessment needs more than a checklist: it needs knowledge of nitrosamine chemistry, manufacturing pathways, raw material quality, degradation mechanisms, analytical limits and current regulatory expectations.
This article explains what “reasonably expected” means in practice, how it changes your risk assessment, and what evidence reviewers look for. It reflects the analytical perspective of ResolveMass Laboratories Inc., a Canadian analytical CRO/CDMO focused on mass spectrometry, biosimilar characterization and nitrosamine testing.
Summary:
- A reasonably expected nitrosamine is one that may plausibly be present or form in a drug substance or drug product, based on its structure, ingredients, manufacturing process, packaging, storage conditions or degradation pathways.
- A FDA Nitrosamine Risk Assessment identifies potential nitrosamines, evaluates how they could form or enter the product, and decides whether confirmatory testing or mitigation is needed.
- A theoretical pathway alone does not prove a nitrosamine is present. Chemical plausibility, process knowledge and analytical data decide the real risk.
- FDA’s guidance covers two classes: small-molecule nitrosamines and nitrosamine drug substance-related impurities (NDSRIs).
- Assessments should cover APIs, excipients, solvents, reagents, recovered materials, equipment and container-closure systems.
- Results are judged against an acceptable intake (AI) limit, and NDSRIs use the Carcinogenic Potency Categorization Approach (CPCA).
- Sensitive, validated mass spectrometry methods from a qualified laboratory support scientifically justified decisions.
1: Why “Reasonably Expected” Drives Your FDA Nitrosamine Risk Assessment
A nitrosamine is “reasonably expected” when your chemistry, process or materials make its formation or presence plausible, and that judgment decides whether your FDA Nitrosamine Risk Assessment stays a documented rationale or leads to confirmatory testing and controls. Certain nitrosamines have shown carcinogenic potential, which is why FDA treats them as a patient-safety and quality priority.
FDA’s guidance, Control of Nitrosamine Impurities in Human Drugs, was revised in September 2024 (Revision 2). It describes root causes of formation and how API and drug product manufacturers should assess, test and control nitrosamines. The phrase “reasonably expected” separates a credible chemical or process risk from a hypothetical possibility with little scientific support. That distinction affects testing decisions, method development, supplier qualification, process changes and regulatory submissions.
This article reflects the analytical perspective of ResolveMass Laboratories Inc., a Canadian analytical CRO/CDMO focused on mass spectrometry, biosimilar characterization and nitrosamine testing.
2: What Is a “Reasonably Expected” Nitrosamine Under FDA Guidance?
A reasonably expected nitrosamine is an impurity with a scientifically plausible route to formation, introduction or persistence in a specific drug substance or product, supported by chemical, manufacturing, formulation, packaging, storage or analytical evidence. FDA does not publish it as a standalone category with a single numerical threshold. It describes the expectation that decides which impurities deserve further evaluation.
A nitrosamine may be reasonably expected when one or more of these factors apply:
- Secondary or tertiary amines, or other nitrogen-containing precursors, are present.
- Nitrite or another plausible nitrosating agent is present under compatible conditions.
- Manufacturing conditions permit nitrosamine formation.
- Starting materials, intermediates, solvents, reagents or excipients are known to carry relevant impurities.
- Recovered materials or shared equipment could cause contamination.
- Degradation or interactions during formulation or shelf life could form the impurity.
- Supplier information, process history, literature or analytical data point to a risk.
Important distinction: a possible pathway is a reason to evaluate, not proof of presence. If the materials never meet, the conditions are incompatible, or reliable evidence shows the pathway is controlled, the risk may be substantially lower.
How does “reasonably expected” differ from a detected nitrosamine?
| Status | Basis | What it means for you |
|---|---|---|
| Detected | Analytical result above the limit of quantitation | Compare with the AI limit and act on any exceedance |
| Reasonably expected | Plausible amine, nitrosating agent and conditions | Confirmatory testing or a documented, evidence-based rationale |
| Theoretical only | No credible route to formation, with evidence | Document the justification; testing may not be needed |
3: Why Does “Reasonably Expected” Matter in FDA Nitrosamine Risk Assessment?
It lets manufacturers focus on credible formation pathways instead of testing indiscriminately for every conceivable nitrosamine. A sound FDA Nitrosamine Risk Assessment answers four questions:
- What could be present? Identify relevant nitrosamines and their precursors.
- How could it form or enter the product? Examine manufacturing, formulation, contamination and degradation pathways.
- How likely is it, and at what level? Weigh process conditions, controls, data and analytical findings.
- What action is justified? Decide on testing, process changes, supplier controls or other mitigation.
The assessment should also explain why certain nitrosamines were selected and why other theoretical possibilities were excluded. Quality reviewers and regulatory submissions rely on that rationale.
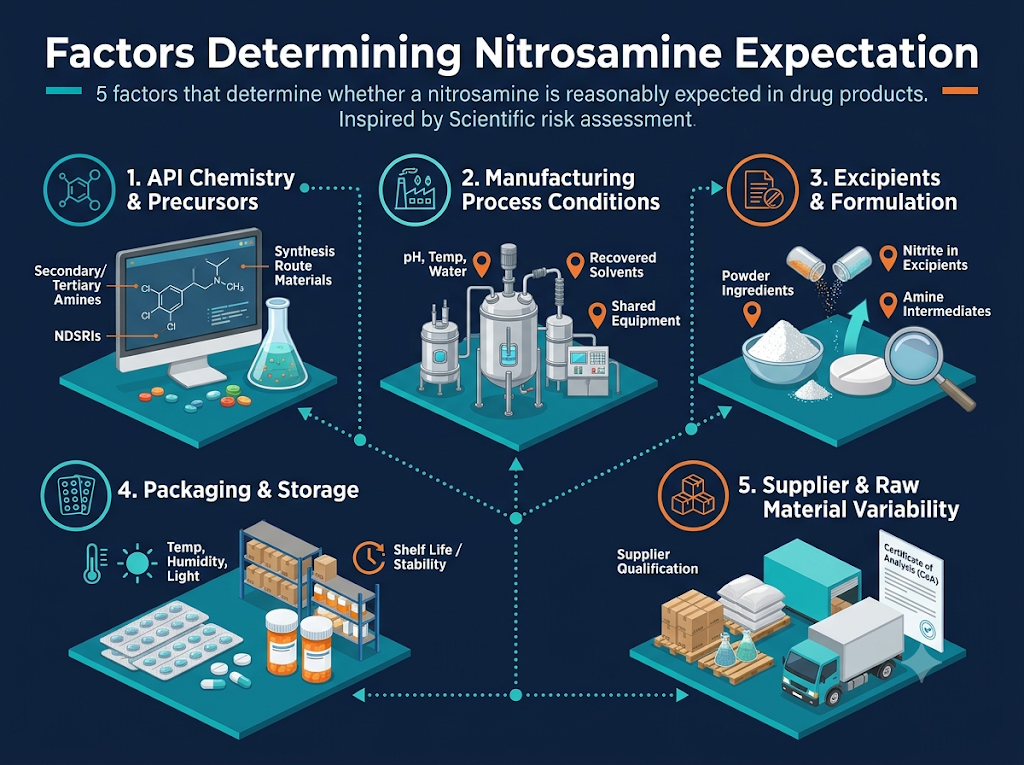
4: Which Factors Determine Whether a Nitrosamine Is Reasonably Expected?
The determination depends on the full product and manufacturing context; one ingredient or process step rarely settles it. For a deeper look at the chemistry, see our guide to nitrosamine formation risk in drug products.
1. API chemistry and nitrosamine precursors
An API’s structure shows whether nitrosamine formation is plausible. Review secondary and tertiary amines, other nitrosatable groups, the synthesis route, starting materials, intermediates, reagents and known impurities. NDSRIs, which are structurally related to the API, need particular attention, and our guide on trace-level NDSRI testing explains how they are approached analytically. Molecule class matters too: see our resources on nitrosamine risk assessment for peptide APIs and a worked case, nitrosamine risk assessment for a proton pump inhibitor. Structural alerts are screening tools, not proof of formation.
2. Manufacturing process conditions
pH, temperature, reaction time, water content, reagent concentration and order of addition all influence formation. Also assess:
- Recovered solvents and reagents
- Recycled process streams
- Cross-contamination and shared equipment cleaning
- Changes in suppliers, materials or manufacturing sites
- Whether impurities can survive purification and crystallization
3. Excipients and drug product formulation
Nitrosamines can form after the API meets excipients. Consider nitrite in excipients, amine-containing ingredients, moisture, pH and interactions during storage, and evaluate formation over shelf life where the formulation justifies it.
4. Packaging and storage conditions
Container-closure systems may contribute contaminants or promote formation when a credible pathway exists. Temperature, humidity, light and storage time affect stability, and well-designed stability studies show whether a suspected nitrosamine increases over time.
5. Supplier quality and raw material variability
Starting materials, solvents, reagents and excipients can vary by supplier and batch. Consider supplier qualification, impurity specifications, certificates of analysis, change notifications and process information, and add targeted testing or tighter supplier controls where a pathway is credible.
5: Where Do Reasonably Expected Nitrosamines Come From?
They come from the API structure, reagents and solvents, quenching steps, vendor-sourced raw materials, excipients, packaging and storage, all of which FDA’s guidance discusses as root causes.
| Source | Typical concern | Evidence to gather |
|---|---|---|
| API structure | Amine that can form an NDSRI | Structure review, degradation and stability data |
| Reagents and solvents | Amines and nitrite in the same or a later step | Process flow diagrams, step-level chemistry |
| Quenching process | Nitrite used to destroy azides, with amines present | Reaction conditions and carryover data |
| Vendor raw materials | Nitrosamines already in supplied material | Supplier declarations backed by test data |
| Recovered solvents | Cross-contamination between processes | Recovery and cleaning records |
| Excipients | Trace nitrite reacting with an amine API | Excipient nitrite levels, formulation pH |
| Packaging and storage | Certain packaging components, heat and humidity | Packaging composition, stability studies |
6: How Does a Reasonably Expected Nitrosamine Change Your FDA Nitrosamine Risk Assessment?
It turns the assessment into a decision: you either show with a validated test that the nitrosamine is absent or below limits, or you justify with evidence why it cannot form. A structured approach improves consistency and traceability. Use this framework as a starting point and adapt it to current FDA guidance.
| Stage | Key activities | Expected outcome |
|---|---|---|
| 1. Product and process mapping | Review API synthesis, formulation, excipients, packaging, storage | Potential sources and pathways identified |
| 2. Precursor evaluation | Examine amines, nitrites, nitrosating agents, relevant impurities | Plausible nitrosamine candidates established |
| 3. Pathway assessment | Evaluate contact between precursors and process conditions | Credible pathways separated from speculation |
| 4. Existing controls review | Examine supplier controls, purification, process limits | Safeguards and gaps identified |
| 5. Risk characterization | Weigh likelihood, potential levels, exposure, uncertainty | Risks needing investigation prioritized |
| 6. Confirmatory testing | Apply suitable analytical methods where warranted | Presence confirmed and quantified, or ruled out |
| 7. Mitigation and verification | Implement controls and verify they work | Risk reduced and ongoing control established |
Document the evidence behind each conclusion. If uncertainty remains, the next step is a proportionate investigation, not an unsupported assumption that the risk is negligible or unacceptable.
What supports a “no risk” conclusion?
- A structure-based review of the API and all intermediates for nitrosatable amines
- Step-by-step process chemistry showing where nitrosating agents could appear
- Data-backed supplier information for key materials, solvents and excipients
- Stability and packaging considerations, with dates and reviewers recorded
- A plan to revisit the assessment whenever a process, supplier or formulation changes
7: When Is Confirmatory Testing Necessary?
Confirmatory testing is needed when the assessment finds a credible potential for nitrosamine presence, or when evidence cannot resolve a significant uncertainty. Design the test around the suspected compound, expected concentration and product matrix. If you are setting up capability for the first time, read our guide to building a nitrosamine testing program from scratch.
Typical analytical techniques:
- LC-MS/MS: sensitive, targeted quantification of known nitrosamines
- LC-HRMS: accurate-mass information for unexpected or related impurities
- GC-MS or GC-MS/MS: suitable for volatile or thermally stable nitrosamines
- Other fit-for-purpose approaches: chosen by analyte, matrix and required sensitivity
Choose the platform by the scientific question, not by assuming one instrument suits every nitrosamine. Result quality also depends on nitrosamine sample preparation and extraction, because matrix effects and contamination can distort trace-level results.
What should a confirmatory method demonstrate?
- Specificity and selectivity
- Sensitivity and an appropriate limit of quantitation
- Accuracy and precision
- Calibration range and linearity
- Recovery from the product matrix
- Robustness and sample stability
- Controls against contamination and carryover
A method that cannot separate the target from matrix interference can mislead, and a result below the detection limit is not automatic proof of absence.
8: How Do Acceptable Intake Limits and CPCA Affect Risk Assessment?
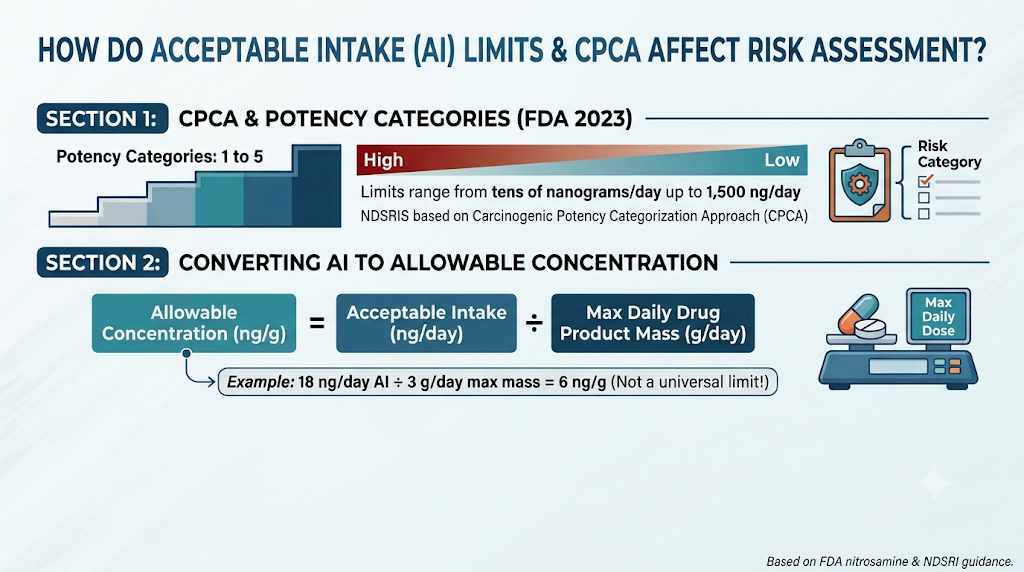
Once a nitrosamine is measured, you compare it with its acceptable intake (AI) limit, which depends on the specific compound and current FDA recommendations. For NDSRIs, FDA’s 2023 guidance on recommended AI limits uses the CPCA to place each impurity in one of five predicted potency categories, with limits ranging from tens of nanograms per day up to 1,500 ng per day.
An AI is expressed per day, so you convert it using the maximum daily dose:
Allowable concentration (ng/g) = Acceptable intake (ng/day) ÷ Maximum daily drug product mass (g/day)
For an API-based concentration in ng/mg, use the daily API dose in mg/day. As an illustration only, an AI of 18 ng/day with a 3 g/day maximum daily product mass gives 6 ng/g. That 18 ng/day figure is not a universal limit.
Timeline for approved products: FDA recommended that confirmatory testing for at-risk products be completed and necessary changes submitted by August 1, 2025. On June 23, 2025, FDA updated its AI limits webpage to say not every approved application could meet that date, allowing more time for changes and progress updates. Because these dates have passed and FDA updates its pages, check the current CDER nitrosamine AI limits page before filing. If no compound-specific AI exists, consult current guidance rather than assigning an arbitrary limit. Products for several markets may face different regulator limits, so confirm the strictest applicable one.
How Does the Assessment Influence Regulatory and Quality Decisions?
It shapes testing, process development, supplier management, stability programs and submissions, and its conclusions should match the evidence. Possible actions include:
- Documenting why a pathway is not scientifically credible
- Running targeted confirmatory testing
- Revising raw material or finished-product specifications
- Strengthening supplier qualification and impurity controls
- Modifying process conditions or evaluating alternative materials or routes
- Strengthening stability monitoring or packaging controls
- Opening corrective and preventive actions when warranted
- Assessing reporting obligations and regulatory submission changes
If a nitrosamine is detected, evaluate its identity, concentration, patient exposure and applicable AI. Detection alone does not make a product unsafe, and a negative result from an insufficiently sensitive method does not prove the risk is absent.
9: Common Gaps in an FDA Nitrosamine Risk Assessment
The most common gap is concluding “no risk” without showing the reasoning.
| Common mistake | Why it matters | Better approach |
|---|---|---|
| Treating every theoretical possibility as confirmed | Unnecessary, poorly prioritized testing | Evaluate plausibility and actual process conditions |
| Assessing only the API | Misses formulation-related formation or contamination | Include excipients, packaging and storage |
| Relying only on certificates of analysis | May miss process-specific trace impurities | Combine supplier documents, process knowledge and targeted testing |
| Using a method with insufficient sensitivity | May miss impurities at relevant exposure | Establish fit-for-purpose sensitivity |
| Applying one AI to every nitrosamine | Limits differ by compound | Verify the current compound-specific recommendation |
| Ignoring supplier or process changes | May introduce new precursors | Reassess when changes occur |
| Not documenting uncertainty | Conclusions are hard to defend | Record assumptions, evidence, limits and follow-up |
How ResolveMass Laboratories Supports Your FDA Nitrosamine Risk Assessment
ResolveMass Laboratories Inc. is a Canadian analytical CRO/CDMO that combines pharmaceutical science expertise with mass spectrometry-based analytical work, including nitrosamine testing. Our scientists support the evidence side of an FDA Nitrosamine Risk Assessment and the testing that follows. Teams working on complex biologics can also review our nitrosamine compliance strategy for biosimilars.
| Need | How ResolveMass helps |
|---|---|
| Identifying plausible nitrosamines | Review of API structure, process chemistry and materials to flag credible pathways |
| Targeted nitrosamine analysis | Appropriate analytical approach for a specific suspected impurity |
| Method development | Sensitive, selective LC-MS/MS methods suited to the analyte and matrix |
| Impurity characterization | High-resolution MS investigation of unexpected or related impurities |
| Data interpretation | Results reviewed against method performance, product composition and AI limits |
The analytical plan is defined after reviewing the suspected nitrosamine, sample type, required reporting limit, maximum daily dose and regulatory criteria. Confirm specific services, validated methods, accreditation and turnaround times directly with ResolveMass before relying on them for a project. Final regulatory conclusions remain the responsibility of the license holder.
References
- Control of Nitrosamine Impurities in Human Drugs, Revision 2 (FDA, September 2024)
- Recommended Acceptable Intake Limits for NDSRIs (FDA, August 2023)
- CDER Nitrosamine Impurity Acceptable Intake Limits (FDA)
- Nitrosamine Impurities in Medications (FDA)
- FDA extends deadline for nitrosamine testing submissions (RAPS, June 2025)
Conclusion: Build a Defensible FDA Nitrosamine Risk Assessment
A reasonably expected nitrosamine is one supported by a credible scientific pathway, not merely one that could theoretically exist, and it decides whether your FDA Nitrosamine Risk Assessment ends in a documented rationale or in validated confirmatory testing. The strongest assessments review the full supply chain, justify every “no risk” conclusion with evidence, use methods sensitive enough for the AI limit, and are revisited whenever data, processes or regulations change.
Frequently Asked Questions:
Manufacturers review the API structure, amine-containing ingredients, nitrites or other relevant nitrosating agents, manufacturing conditions, excipients, packaging, and storage conditions. Chemical plausibility and process-specific evidence help determine whether a formation pathway warrants further investigation.
No. A potential formation pathway indicates a possible risk, not confirmed contamination. Additional scientific evaluation or confirmatory analytical testing may be necessary to establish whether the nitrosamine is present and at what concentration.
A conventional small-molecule nitrosamine impurity may originate from reagents, solvents, or process-related contaminants. A nitrosamine drug substance-related impurity (NDSRI) is structurally related to the active pharmaceutical ingredient and may form through reactions involving the drug substance or related precursors.
Confirmatory testing is generally appropriate when a risk assessment identifies a credible potential for nitrosamine presence or when significant uncertainty remains that cannot be resolved through existing evidence. The testing strategy should address the suspected compound, product matrix, and relevant reporting or quantification requirements.
An acceptable intake (AI) is a compound-specific or otherwise scientifically established amount used to evaluate potential patient exposure. The applicable limit depends on the nitrosamine and current regulatory recommendations; there is no single universal AI limit for all nitrosamines.
A simplified calculation is:
Acceptable concentration (ng/g) = Acceptable intake (ng/day) ÷ Maximum daily drug product mass (g/day)
The calculation must use consistent units and the appropriate daily product quantity. The result should then be evaluated against the applicable regulatory requirements and the actual or estimated patient exposure.
Reference
- Pardeshi N, Satapara V, Patel K. Nitrosamine contamination in pharmaceuticals: Regulatory perspectives and control strategies of USFDA, EMA & HC. International Journal Of Drug Regulatory Affairs. 2024;12(1):1-8.https://indianjournals.com/api/article-view/ijdra-12-1-001
- Umrethwala A, Mutalik R, Sharma S. Nitrosamine Impurities: Critical Insights into Chemistry, Analytical Tools, and Worldwide Regulatory Requirements. InAnnales Pharmaceutiques Françaises 2026 Apr 4. Elsevier Masson.https://www.sciencedirect.com/science/article/pii/S0003450926000441
- Johnson GE, Dobo K, Gollapudi B, Harvey J, Kenny J, Kenyon M, Lynch A, Minocherhomji S, Nicolette J, Thybaud V, Wheeldon R. Permitted daily exposure limits for noteworthy N‐nitrosamines. Environmental and Molecular Mutagenesis. 2021 Jun;62(5):293-305.https://onlinelibrary.wiley.com/doi/abs/10.1002/em.22446
- Charoo NA, Dharani S, Khan MA, Rahman Z. Nitroso impurities in drug products: an overview of risk assessment, regulatory milieu, and control strategy. AAPS PharmSciTech. 2023 Feb 9;24(2):60.https://link.springer.com/article/10.1208/s12249-023-02523-w
- BHENDE S, PHATAK A, CHAUDHARI P. Risk-Based Determination of Analytical Sensitivity (LOQ) for Nitrosamine Impurities Using Acceptable Intake Limits.https://www.academia.edu/download/134656412/60796.pdf
About the Author